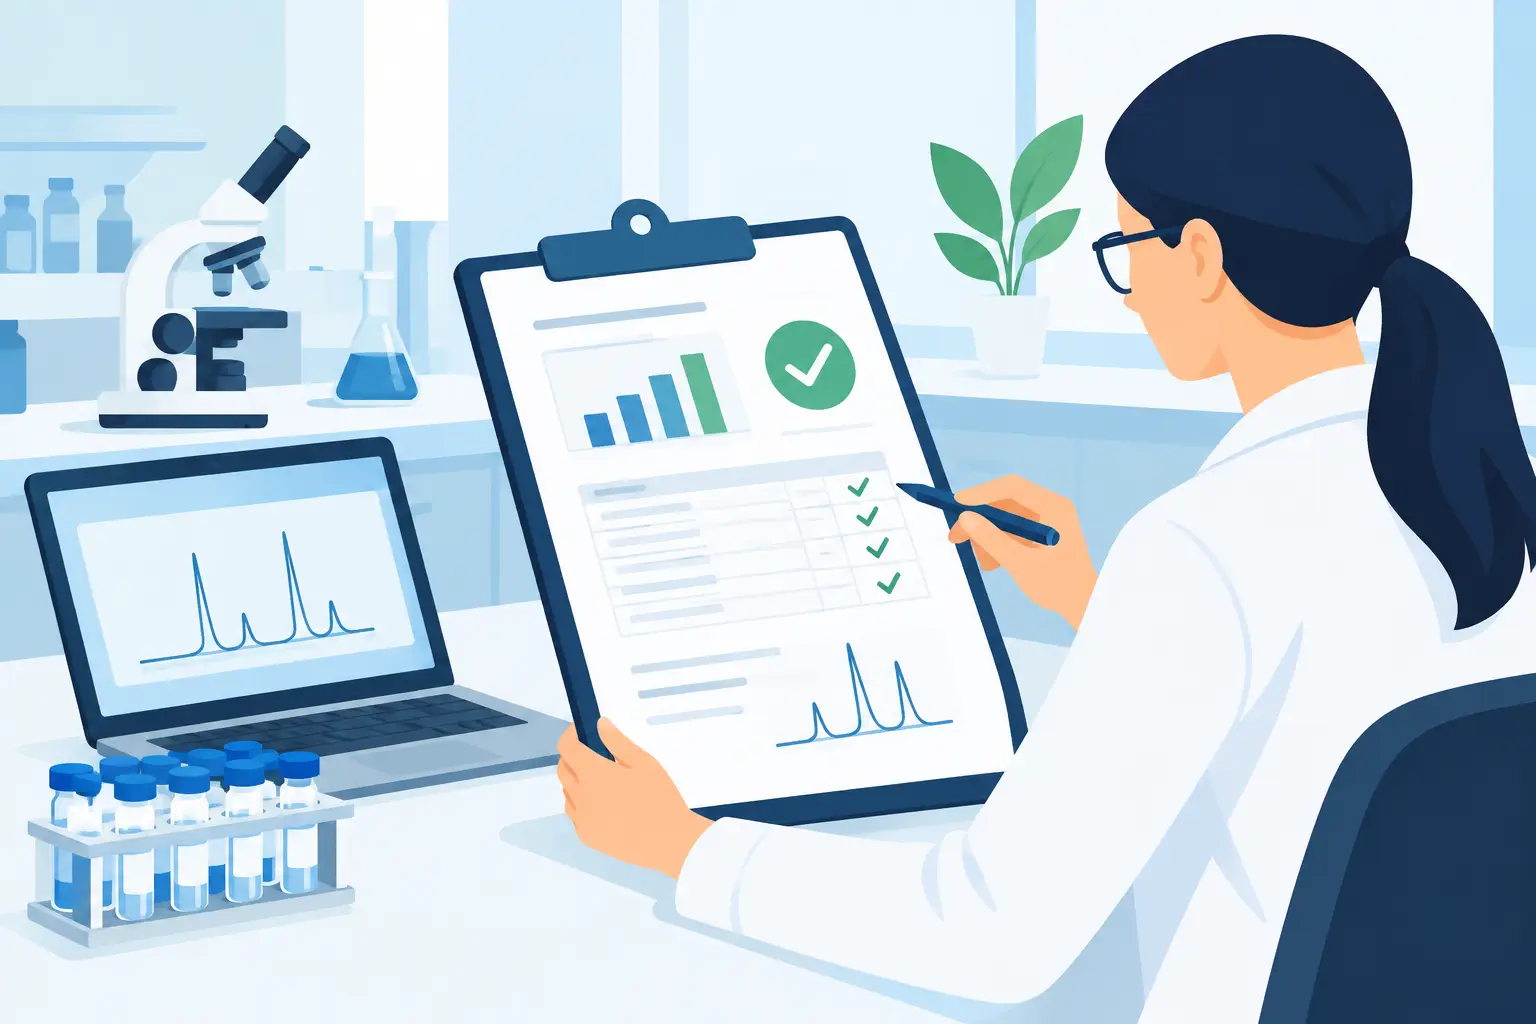
A report can look technical and still leave the central research question unanswered: does this document clearly support the specific batch in hand? An assay report format review is the disciplined process of answering that question before a material is entered into a research workflow. It examines more than a passing result. It checks whether the record connects the sample, analytical method, result, specification, and batch identity in a controlled, readable chain.
For research peptide purchasers, format matters because documentation is part of traceability. A percentage without a sample identifier is not batch verification. A chromatogram without a method reference is difficult to interpret. A certificate that identifies a product but not the lot may establish a general claim, not evidence for the material received.
What an Assay Report Must Establish
An assay report is not interchangeable with a marketing specification, a generic product data sheet, or a statement that testing occurred. Its purpose is to document an analytical result for a defined sample under a defined method. Depending on the laboratory and material, the reported assay may address content, purity, identity, concentration, potency, or another measured characteristic. The word assay alone does not tell the full story.
A useful report establishes five connected facts. It identifies the material tested, identifies the lot or batch, states the analytical method, reports the measured result with appropriate units, and shows how that result is evaluated against an acceptance criterion or specification. If one of these elements is absent, the reader must decide whether the remaining evidence is sufficient for the intended research use.
That standard should be applied proportionally. A preliminary supplier screen may begin with a high-level certificate. Material intended for controlled, repeatable laboratory work requires stronger lot-level documentation, including dates, method details, and review controls. The correct threshold depends on the study, but the need for an unbroken documentation path does not.
Assay Report Format Review: Start With Document Control
Begin with the information that ties the report to a physical material. The product name should be specific enough to distinguish the item from related compounds, salts, concentrations, or presentations. The lot or batch number should be visible and should match the identifier used on the product label, packaging record, or supplier verification system.
A report date is useful, but it is not a substitute for a test date. When both appear, they answer different questions. The test or analysis date indicates when the work was performed. The issue date indicates when the document was released. A retest date or expiration date, where applicable, should be interpreted within the supplier's stability and quality system rather than treated as proof of current performance by itself.
The document should also show a report number, sample number, or another controlled identifier. This creates an audit path if a question arises later. In a well-organized quality documentation system, the batch record, COA, and verification result are not isolated files. They are records that can be reconciled to the same lot.
Verify the sample description
The sample description deserves close attention. A report may refer to a received sample, a retained sample, a composite, or a finished material. These are not automatically equivalent. A composite result can provide useful process information, but it may not establish the result for each individual unit. Likewise, a result generated on an intermediate may not apply to the final packaged material without clear traceability.
For peptide materials, confirm details that can change interpretation, such as the stated sequence or identifier, salt form, mass basis, and presentation. A report that uses an internal code can still be acceptable when that code is clearly mapped to the purchased product and lot. If the mapping is not visible, the report is incomplete from the purchaser's perspective.
Read specifications as decision criteria
A result has limited meaning without context. The report should state the acceptance criterion beside the result or provide an unambiguous reference to the governing specification. For example, a purity result of 98.4% is meaningful only when the method and required threshold are known.
Pay attention to inequality signs, decimal places, and units. A result reported as greater than 98% is different from a numerical result of 98.0%, particularly when the release limit is close. Similarly, assay values expressed on an as-is basis, anhydrous basis, or salt-free basis cannot be compared casually. The format should make the basis clear.
Evaluate the Method Before Interpreting the Result
A clean report format does not validate an unsuitable method. The method must be capable of addressing the attribute claimed. High-performance liquid chromatography may be used to evaluate purity profiles, while mass spectrometry can support molecular identity. Neither result should be presented as if it answers every quality question.
A chromatographic purity percentage, for instance, does not independently confirm sequence, biological activity, sterility, endotoxin status, residual solvents, or water content. It reports what the stated method measures under the stated conditions. That limitation is not a defect. It is the normal boundary of analytical evidence.
Method information should be sufficient to understand the test at an appropriate level. Depending on the documentation model, this may include the analytical technique, method identifier, detector, column chemistry, wavelength, mobile phases, run conditions, or a controlled reference to an internal validated method. External purchasers do not always need proprietary procedural details, but they should be able to identify what was done and distinguish one test from another.
Review supporting data with restraint
Supporting data can strengthen a report when it is legible and connected to the sample. Chromatograms should have identifiable axes, retention times, integration details where relevant, and a sample or injection reference. Mass spectra should identify the expected mass relationship and the specimen tested. Screenshots without identifiers or clipped graphs without scale provide less assurance than complete, controlled attachments.
Still, more pages do not automatically mean better evidence. An extensive packet may contain raw-looking data while omitting the batch number, acceptance criterion, or reviewer approval. The purpose of review is not to reward document volume. It is to determine whether the evidence is traceable, interpretable, and fit for the intended decision.
Look for Review Controls and Exceptions
A released quality record normally includes an authorized review or approval indicator. This might be a signature, electronic approval, quality-unit release statement, or controlled certificate status. The exact format varies by laboratory, but the report should indicate that the result was reviewed rather than simply generated by an instrument.
Exceptions require direct attention. Terms such as deviation, investigation, out-of-specification, retest, amended report, or superseded certificate should not be ignored because the final line says pass. A documented retest can be appropriate when it follows defined procedures. It also raises questions: Which result governed release? Why was retesting required? Does the amended report clearly preserve the original record and explain the change?
Version control matters here. If multiple reports exist for one lot, determine which is current and whether the later document replaces or supplements the earlier one. A secure batch verification process should make current status clear without erasing the documentation trail.
Build a Practical Review Workflow
A consistent workflow reduces the risk of accepting a document based on appearance alone. First, match the product and lot number to the item received. Next, confirm that the report identifies the sample and provides a controlled document reference. Then review the method, result, units, and specification together rather than as separate fields.
After that, evaluate whether the reported test supports the specific attribute you need for the research plan. If purity is the decision point, confirm that the method and purity result are clear. If identity is essential, determine whether identity-specific evidence is provided. If the research requires additional attributes, such as bioburden or residual solvent limits, look for separate testing records instead of assuming an assay result covers them.
Finally, retain the record with the lot information used in your internal files. For digitally managed purchasing, a supplier's COA Registry or batch verification resource can simplify retrieval, but the purchaser should still reconcile the document to the material received. Axis Direct Labs organizes batch-level verification and comprehensive quality documentation around this traceability requirement.
A well-formatted assay report does not eliminate the need for scientific judgment. It gives that judgment a controlled foundation: a specific material, a stated method, a defined result, and a record that can be checked again when the research demands it.